PRODUCT NEWS · GLOBAL SUBMISSION PORTAL
Manage electronic instructions for use centrally, control approvals and make them digitally accessible.
With the new eIFU module, Europe IT Consulting expands the Global Submission Portal (GSP) with an additional solution for the digital management of instructions for use. The extension is available as an optional module and is activated individually for each customer after purchase.
Basel, 19 November 2026 – Europe IT Consulting GmbH is expanding its Global Submission Portal (GSP) with a new module for electronic Instructions for Use (eIFU). The solution provides medical device manufacturers with an integrated way to manage product-related instructions for use digitally, maintain traceable document versions and make approved information available to users.
The new eIFU module complements the GSP’s existing capabilities for UDI data management and regulatory data submission. Europe IT Consulting is thereby extending the platform into another area of digital product documentation, without requiring manufacturers to introduce a separate software environment.
Electronic Instructions for Use Are Becoming Increasingly Important
Medical device manufacturers face the challenge of reliably managing instructions for use across numerous products, variants and language versions. Changes must be traceable, documents must pass through internal review and approval processes, and the resulting information must be made available to the intended users.
At the same time, the European regulatory framework for electronic instructions for use continues to evolve. Commission Implementing Regulation (EU) 2025/1234 expanded the existing provisions. Under the conditions established by law, instructions for use for medical devices intended for professional users may be provided electronically.
For manufacturers, this means that structured document management is becoming increasingly important alongside digital publication.
eIFU Internet Addresses in EUDAMED and swissdamed: What Manufacturers Need to Know
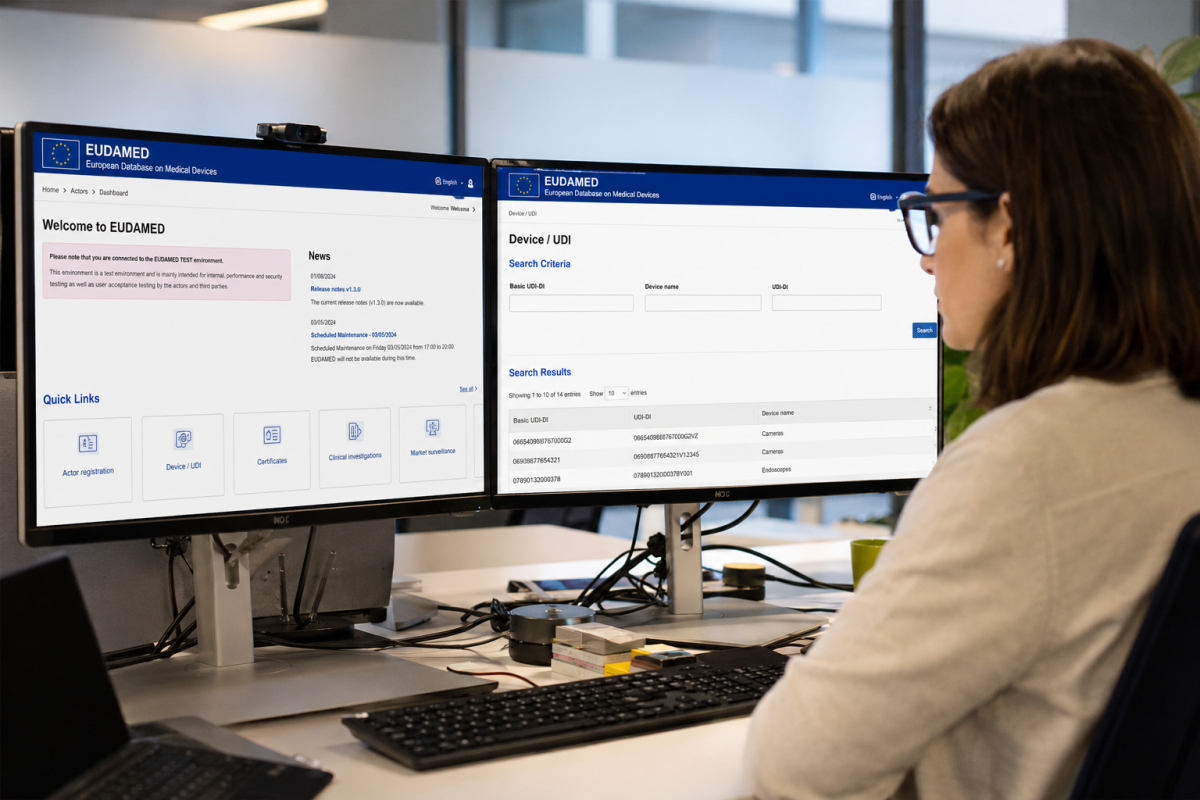
With mandatory UDI device registration, the digital provision of instructions for use is gaining additional regulatory significance. It is important to distinguish between the instructions themselves and their internet address: There is no general obligation to upload IFU PDF files to EUDAMED or swissdamed. However, when electronic instructions for use are provided instead of paper versions in accordance with the applicable rules, the corresponding internet address may be mandatory in the respective UDI device record.
EUDAMED: For electronic instructions for use governed by Commission Implementing Regulation (EU) 2021/2226, as amended, Article 7(3) requires manufacturers to provide the relevant internet address in the UDI database. Mandatory use of the UDI/Devices module began on 28 May 2026. For certain devices placed on the market before that date which remain subject to registration, the European Commission provides for a transition period until 28 November 2026.
swissdamed: Swissmedic also requires the internet address at which the electronic instructions for use can be accessed when registering medical devices provided with an eIFU. Mandatory Swiss device registration began on 1 July 2026; for the relevant devices, a transition period generally applies until 31 December 2026.
The expanded EU eIFU rules generally apply to medical devices under the MDR regardless of risk class, provided they are intended for professional users and the legal conditions are met. There is no general obligation to replace paper instructions for use with eIFUs. In vitro diagnostic medical devices under the IVDR are subject to separate provisions and cannot automatically be treated in the same way as MDR devices under the eIFU rules.
For manufacturers using eIFUs, this makes not only structured document management important but also the reliable provision of a stable, accessible eIFU internet address. Europe IT Consulting’s new eIFU module supports manufacturers in the structured management and digital provision of electronic instructions for use, complementing the Global Submission Portal’s existing UDI capabilities.
Official sources: EU Implementing Regulation 2025/1234 · European Commission: UDI/Devices · Swissmedic: swissdamed FAQ
The New eIFU Module in the Global Submission Portal
The new module from Europe IT Consulting supports the management and publication of electronic instructions for use within the existing Global Submission Portal. Its purpose is to bring relevant documentation processes together in a common environment and simplify the organization of eIFU documents for manufacturers.
Instructions for use can be managed in a structured manner, assigned to the corresponding products and processed through controlled review and approval workflows before publication. Different language versions and document revisions can also be organized with traceability.
An important advantage is the connection with existing UDI product information. This enables manufacturers to use their available product data when assigning electronic instructions for use to the corresponding devices.
Key Features at a Glance
- Centralized management: Organize and maintain electronic instructions for use in a structured environment.
- Product assignment: Associate eIFU documents with the corresponding UDI-DI product information.
- Controlled approvals: Support internal review and approval processes before publication.
- Version management: Maintain traceable records of changes and document revisions.
- Multilingual support: Manage and provide instructions for use in different language versions.
- Public eIFU portal: Make approved and published documents digitally accessible to users.
Digital Access to Instructions for Use
Through a publicly accessible eIFU portal, users can search for published instructions for use, view them and download them as PDF documents. Search options based on document and product information, together with the available language versions, make it easier to access the required documentation.
The solution therefore supports both the manufacturer’s internal documentation processes and the external provision of product-related information to healthcare professionals and other intended users.
eIFU and UDI Data Management on One Platform
The Global Submission Portal already supports medical device manufacturers in preparing, validating and electronically submitting UDI data to regulatory databases such as EUDAMED, swissdamed, FDA GUDID and TGA AusUDID. With the new eIFU extension, Europe IT Consulting adds digital management and publication of electronic instructions for use to these existing capabilities.
The shared platform provides an opportunity to better coordinate product information and the associated documentation. For Regulatory Affairs, Quality Management and Technical Documentation teams in particular, this creates opportunities to organize their activities more centrally.
Optional Add-On Module for GSP Customers
The new eIFU module is an additional extension of the Global Submission Portal and is not automatically included in the standard activated functionality. Interested customers can purchase the module separately. It is then activated individually for the respective customer.
Which Manufacturers Can Benefit from the eIFU Module?
The extension is designed for medical device manufacturers seeking to manage electronic instructions for use in a structured manner, maintain multiple product or language versions, or provide users with convenient digital access to documentation.
Existing GSP customers can selectively expand their current functionality. New customers can include eIFU management when configuring the GSP capabilities required for their organization.
Discover the New eIFU Module
Would you like to learn more about the capabilities of the new eIFU module and how it can be integrated into the Global Submission Portal? Europe IT Consulting will be pleased to provide information about the available functionality, booking options and potential applications for your company.
Regulatory note: The electronic provision of instructions for use is subject to the applicable legal requirements and conditions. Responsibility for compliance remains with the medical device manufacturer.
Legal basis: Commission Implementing Regulation (EU) 2021/2226, as amended by Commission Implementing Regulation (EU) 2025/1234.






Related Posts