Understanding EUDAMED · Europe IT Consulting
EUDAMED: structure, modules and how they connect
EUDAMED is the European database on medical devices. Operated by the European Commission, the system connects information on actors, devices, certificates, studies, vigilance and market surveillance. The UDI/Device module is therefore one part of the overall system, rather than the entire database.
This distinction matters in practice for manufacturers: registering the company, maintaining UDI data and managing information from a notified body are different tasks. A data process must therefore provide the correct values and account for the correct roles and relationships.
Here you will find the structure of the six modules, their current status and what they mean for your UDI data work. If you are already looking for a specific submission method, the EUDAMED decision guide takes you directly to manual entry, XML and M2M.
Sources checked: · For Regulatory Affairs, quality management and IT
The six EUDAMED modules at a glance
The table distinguishes purpose, participating roles and usage status. Here, “mandatory” describes the module status. It does not mean that every manufacturer must use every module themselves or that the same deadline applies to all existing records.
| Module | Purpose and participants | Status |
|---|---|---|
| Actor Registration Actors and Actor ID/SRN |
Identity and role of the organisations involved Economic operators and competent authorities |
Mandatory since 28 May 2026 |
| UDI/Device Registration Identification and device data |
UDI and device data in the European context Primarily manufacturers; roles depend on the registration |
Mandatory since 28 May 2026 |
| Notified Bodies & Certificates Notified bodies and certificates |
Certificate information and status Notified bodies; linked actor and device data |
Mandatory since 28 May 2026 |
| Clinical Investigations & Performance Studies Clinical investigations and performance studies |
Study-related applications, notifications and information Sponsors and responsible bodies |
Still under analysis/development |
| Vigilance & Post-Market Surveillance Vigilance and post-market surveillance |
Safety reports and post-market information Manufacturers and competent authorities |
Still under development |
| Market Surveillance Market surveillance |
Information from market surveillance by authorities Competent authorities and the European Commission |
Mandatory since 28 May 2026 |
Current module status · European Commission: EUDAMED overview
Check functionality and mandatory use separately. An available module may contain different functions and access rights. For a particular activity, the applicable transitional rules and current procedure description must also be considered.
How are actors, devices and certificates connected?
For UDI data work, a simple set of questions helps: who is the responsible organisation? Which device group and devices are being described? Which certificate information, if applicable, is linked to them?
- Actor · Who is involved?
- The registered organisation provides the reference for its data and users.
- Device data · What is described?
- Basic UDI-DI, UDI-DI and the relevant attributes represent the device structure.
- Certificate relationship · Which information belongs here?
- In the applicable procedure, certificates are recorded with the appropriate actor and device references.
Not a linear six-step process: A device does not automatically pass through all six modules as successive registration stages. Activities differ according to role and purpose.
European Commission: Actor Registration · European Commission: Notified Bodies & Certificates

What each module does
The following sections explain the role of each area and provide practical guidance for preparing your data and responsibilities.
 Module 01
Actor Registration
Actors and Actor ID/SRN
Manufacturers, authorised representatives, importers and system/procedure pack producers, among others, register in the Actor module. The competent national authority reviews the registration request; once approved, the Actor ID/SRN is generated in EUDAMED.
Important when clarifying roles: A Competent Authority (CA) is the responsible authority. A Notified Body (NB) is a notified body. Actor registration should not be understood as a separate SRN application to a notified body.
Organisations and users also differ: anyone working on behalf of a registered actor needs the appropriate user access.
European Commission: Actor Registration
For your preparation: Keep actor data, user responsibilities and production identifiers consistent. This makes subsequent data relationships clearly traceable.
 Module 02
UDI/Device Registration
Identification and device data
The UDI/Device module associates identifiers and device information with the registered manufacturer. The European Commission provides different datasets for MDR, IVDR and legacy devices. EMDN nomenclature is also used to categorise devices.
European Commission: UDI/Device Registration
For standard MDR/IVDR registrations, distinguish Basic UDI-DI from UDI-DI: the Basic UDI-DI identifies a device group; the UDI-DI identifies a specific device model. Name, model, characteristics and labelling information belong at their designated data level.
EUDAMED help: Basic UDI-DI · EUDAMED help: UDI-DI identification data
For your preparation: Do not simply copy every field from an arbitrary template. Define the legal framework, device relationships and a reliable source for each data field. More about the EUDAMED UDI/Device module.
 Module 03
Notified Bodies & Certificates
Notified bodies and certificates
Notified bodies record certificate information, changes and status here. Referenced actors and Basic UDI-DIs must already be registered for the corresponding relationships. Certificate information is publicly accessible; that does not grant the public editing rights.
European Commission: Notified Bodies & Certificates
Registration does not replace conformity assessment. Whether and to what extent a notified body is involved depends on the applicable procedure. Not every medical device automatically requires an “approval certificate”. A Basic UDI-DI likewise does not automatically establish a fixed one-to-one relationship with exactly one certificate.
Conformity assessment: MDR, Article 52 · MDR: consolidated version of 19 July 2026 · European Commission: Notified Bodies & Certificates
For your preparation: Keep identifiers, device documentation and existing certificate references consistent. Clarify responsibilities with the notified body involved.
 Module 04
Clinical Investigations & Performance Studies
Clinical investigations and performance studies
CI/PS refers to clinical investigations of medical devices and performance studies of in vitro diagnostic devices. The area covers studies and their associated information. It is not simply equivalent to clinical evaluation and PMCF.
European Commission: clinical investigations and performance studies
While the relevant EUDAMED area is not yet available, the procedures applicable to each activity must be followed. For example, the Commission describes publication of clinical investigation reports and summaries outside this EUDAMED module.
European Commission: CI/PS reports
For your planning: Do not equate study-related obligations with the development status of a software feature. Check the specific activity and responsible body; a module name alone is not a submission guide.
 Module 05
Vigilance & Post-Market Surveillance
Vigilance and post-market surveillance
The area provided for in legislation includes reports of serious incidents, field safety corrective actions, trend reports and periodic safety update reports. It is separate from registration of UDI master data.
MDR, Article 92 · MDR: consolidated version of 19 July 2026
For your planning: Organise a safety reporting process separately from routine device data maintenance. The status of a new EUDAMED module does not remove existing obligations or the applicable reporting routes. This page does not provide instructions for a specific incident report.
 Module 06
Market Surveillance
Market surveillance
Market Surveillance concerns surveillance by authorities. The legal framework provides for summaries of surveillance results, inspection reports and information on risks or non-conformities, among other things.
MDR, Article 100 · MDR: consolidated version of 19 July 2026
For context: This area is not another manufacturer form for routine UDI reporting. Distinguish the authorities’ market surveillance from your own post-market surveillance.
Public searches are different from editing access
View information
The public EUDAMED website lets users search information released there, such as published actor, device and certificate information. It is not an openly accessible editor for manufacturer data.
Work for an actor
In the authorised area, the actor and user profile determine the available activities. Having an Actor ID/SRN does not replace the required user access.
European Commission: EUDAMED overview · European Commission: Actor Registration · European Commission: Notified Bodies & Certificates
Which individual information is visible depends on the module, record and release status. A blanket statement that “all information is accessible to everyone” would therefore be misleading. For module areas that are not yet available, distinguish the planned functionality from current access.
What does the structure mean for your UDI data?
The real work often starts before EUDAMED: information is held in Excel, SAP or other structured sources. It needs to become a traceable record. Source, solution, transfer method and operating model are four separate decisions.
DATA SOURCE
Excel, SAP, ERP or another structured data source.
EUROPE IT SOLUTION
GSP, GUDI or an agreed XML project process.
TRANSFER METHOD
Manual entry, manual XML upload or M2M/Data Exchange.
OPERATING MODEL
A one-off project or recurring data maintenance and submission.
EUDAMED help distinguishes manual entry, XML upload and M2M. XML files can be generated automatically, but uploading them through the relevant interface remains manual. M2M is the technical route for exchanging data between connected systems and EUDAMED.
EUDAMED help: data entry and transfer methods
Europe IT: turning data into a manageable submission process
In the Global Submission Portal the customer uploads the completed Excel template. After automatic validation, and with no remaining validation errors, the customer initiates the transfer. GSP uses M2M for EUDAMED.
With GUDI in SAP the customer transfers directly from GUDI using M2M. Submission status and authority feedback support processing. The customer controls data corrections and resubmissions.
In an agreed XML project, Europe IT validates the customer data, then generates XML files and uploads them manually to EUDAMED. These services concern the agreed data process; they do not imply a blanket service covering every function of all EUDAMED modules.
Which transfer method fits your data? · Interpret feedback and validation errors
Device relationships and structured source data
Clear device relationships and a traceable template make data preparation easier. The older examples from the Europe IT page below illustrate the principle; they are not a complete, current EUDAMED field specification.
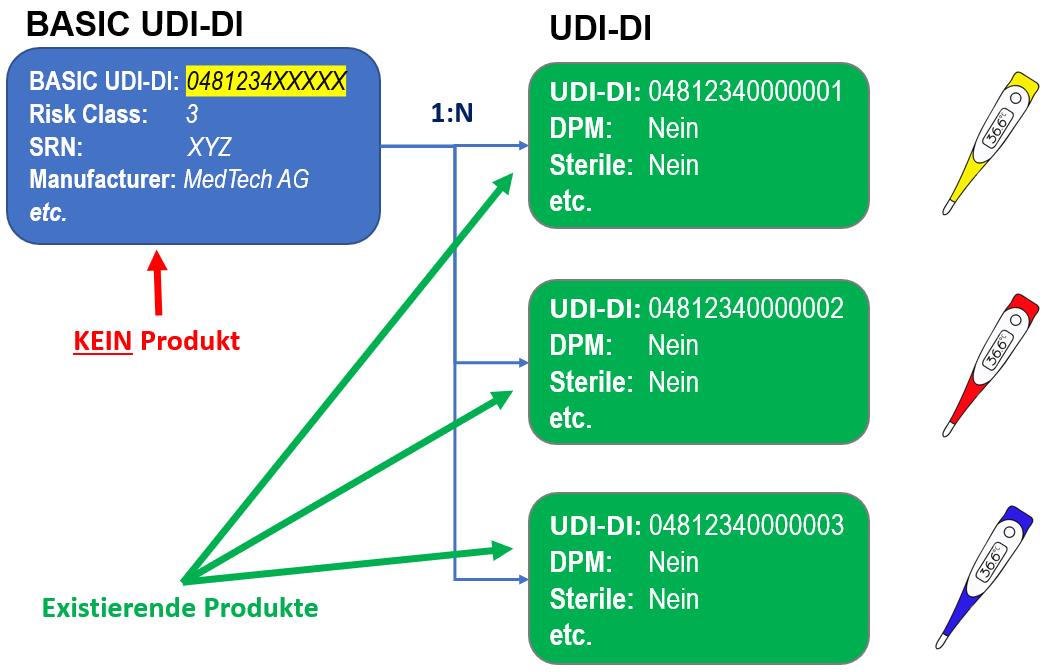
Example: one Basic UDI-DI and multiple UDI-DIs
The diagram shows a one-to-many relationship. The identifiers, risk class and device characteristics shown are illustrative values, not identifier assignment, classification or labelling requirements for the devices pictured.
Example views of the Excel data structure
The existing screenshots show separate data areas and structured columns. For a project, use the current template supplied for that purpose; do not recreate the field list from these images.


Frequently asked questions about EUDAMED structure
Is EUDAMED only a UDI database?
No. UDI/Device is one of six modules. The overall system also covers actors, certificates, clinical investigations and performance studies, vigilance and market surveillance. View the module overview.
Does a manufacturer have to operate all six modules?
No. Responsibilities depend on the role and activity. Entering device data, a notified body’s activities and market surveillance by authorities are different tasks. Responsibilities in detail.
Does a publicly visible record automatically mean device approval?
No. Database registration and the required conformity assessment are different matters. Visibility alone does not provide blanket evidence that every regulatory requirement has been met. Certificate relationships and conformity assessment.
Are XML and M2M separate EUDAMED modules?
No. XML upload and M2M/Data Exchange are transfer methods. They describe how data is submitted; modules describe the subject area. Excel is likewise a possible data source, not an EUDAMED module. Data source, solution and transfer method.
Where can I find help with my specific registration?
For role and registration questions, use the EUDAMED FAQ. For a specific response, consult the EUDAMED error library . This structure page explains the overall context and does not replace instructions for every individual activity.
Related topics
- EUDAMED UDI/Device module: data and structure and UDI/MDR/IVDR terminology.
- MDR/IVDR Excel template and Europe IT solution for EUDAMED.
- MDR terminology and manufacturer obligations (German) and MDR/IVDR timeline and changes – check the source date for any specific deadline.
- Europe IT solution for FDA/GUDID as a separate authority topic.
- Official EUDAMED links and tools and UDI guide.
Sources and currency
Primary sources are linked directly in the relevant sections. Checked on 3 September 2026: current Commission pages on module status and the passages of the consolidated MDR of 19 July 2026 relevant to these statements. Binding legal acts govern legal questions; consolidated versions serve documentation purposes.
Module status, transitional rules and technical features may change. This page explains the structure, rather than providing a complete assessment of the obligations for an individual device. Publishing data in EUDAMED or using a Europe IT solution does not automatically guarantee full regulatory compliance.
