
EUROPE IT · EXPERT KNOWLEDGE FROM A TO Z
UDI, MDR and IVDR glossary: putting technical terms into context
What does the term mean—and what role does it play in your data process?
This glossary explains identifiers, authorities, data formats, roles and reporting terms in the field of Unique Device Identification. Short definitions are complemented by practical distinctions: Basic UDI-DI is not UDI-DI, a technical acknowledgement is not automatically an acceptance, and M2M is a transmission route, not a standalone data-management product.
The entries provide professional orientation. For regulatory decisions, the current requirements of the responsible authority and the specific use case are decisive. Further Europe IT pages provide more detail on each task.
Frequently searched distinctions
Basic UDI-DIMaster UDI-DIM2M – Machine-to-MachineValidationACK / AcknowledgmentGSP – Global Submission PortalGUDI – Global UDI SAP Add-on
A
ACK / Acknowledgment
An acknowledgment is an electronic response to a submission. Receipt, forwarding and substantive processing may be confirmed separately.
The current FDA eMDR FAQ describes four files: ACK1 and ACK2 concern transport, while ACK3 and ACK4 contain the same substantive eMDR processing result in HTML and XML respectively. A receipt ACK alone does not prove substantive acceptance.
Adverse Event Reporting
Adverse Event Reporting means recording, assessing and, where applicable, reporting adverse events or product problems.
The obligation, reporting route and deadline depend on the market, product and role. Product registration in a UDI database is a different task from an incident report.
AIDC – Automatic Identification and Data Capture
AIDC refers to technologies for automatic identification and data capture, such as reading a barcode.
In the UDI context, distinguish the machine-readable carrier, its data content and the human-readable representation.
AIMDD – Active Implantable Medical Devices Directive
AIMDD denotes the former European Directive 90/385/EEC for active implantable medical devices.
For products that previously complied with a directive, the applicable MDR transitional rules must be checked. The abbreviation is not a standalone current database.
AS2 – Applicability Statement 2
AS2 is a standard for the secure exchange of business data over HTTP. Its specification is described in RFC 4130.
The transport protocol does not automatically determine whether the payload contains UDI product data or vigilance reports. An AS2 process must match the respective recipient.
AS4 – Applicability Statement 4
AS4 is a profile for exchanging business messages based on ebMS 3.0.
AS4 is neither an Excel format nor a product name. For an authority connection, the concrete profile, permissions and operating process must be defined.
Audit Trail
An audit trail makes relevant changes or actions traceable in context, for example the user, time, previous value and new value.
A change log is not the same as an authority acknowledgement. Which events are available and how long they are retained depends on the system and process.
AusUDID
AusUDID stands for Australian Unique Device Identification Database, the TGA’s UDI database.
An AusUDID submission must be distinguished from UDI labelling and any required ARTG registration. Sponsor-related assignments are part of the Australian data process.
Authorised Representative
English designation for the authorised representative in the relevant regulatory context.
In EU medical-device law, this is a defined economic-operator role, not automatically every external consultant. See also Authorised Representative.
B
Basic UDI-DI
The Basic UDI-DI is an overarching regulatory identifier in the European UDI system and a key in the associated documentation.
It is not the UDI-DI applied to an individual product and is not an interchangeable synonym for Master UDI-DI. The grouping must meet the applicable criteria.
Authorised Representative under the MDR
An authorised representative takes on defined tasks for a manufacturer outside the EU on the basis of a written mandate.
The central obligations are set out in Article 11 of the MDR, not in a “Chapter 11”. The products and tasks covered by the mandate must be unambiguous.
Primary source / further guidance ↗
WeiterlesenBUDI
BUDI is commonly used in practice as a short form for Basic UDI-DI.
An abbreviated column name should be explained in data models and work instructions. BUDI is not an additional identifier alongside the Basic UDI-DI.
Bulk Upload
Bulk Upload means the bundled processing or submission of multiple records.
Bulk describes the quantity or bundling, not necessarily a particular transport route. A file-based upload and an automated M2M transmission can be different processes.
C
CA – Competent Authority
Competent Authority means the responsible authority. It performs the supervisory and administrative tasks assigned to it by the applicable legal framework.
A competent authority is not the same as a Notified Body. European conformity assessment also does not automatically amount to a single authority approval of a product.
CHRN – Swiss Single Registration Number
CHRN is the Swiss registration number for economic operators in the relevant Swissmedic context.
It does not identify an individual product record and replaces neither the UDI-DI nor any required product registration. Do not mix up CHRN and the European SRN.
CND – Classificazione Nazionale dei Dispositivi medici
CND is the Italian medical-device nomenclature selected as the basis for the European EMDN.
CND and EMDN are not identical without checking the version and application. In EUDAMED, the designated EMDN is relevant.
Code 128
Code 128 is a linear barcode symbology with high data density. GS1-128 is an application built on this symbology within the GS1 system.
A symbology alone does not make a barcode UDI-compliant. Identifier, syntax, print quality and use must work together.
CSDT – Common Submission Dossier Template
CSDT denotes a common structure for technical medical-device submission dossiers in the ASEAN context.
A dossier template structures regulatory documentation. It is neither a UDI-DI nor automatically the technical format of a UDI database.
D
DataMatrix
DataMatrix is a two-dimensional data carrier. GS1 DataMatrix uses this symbol type with the rules of the GS1 system.
Not every DataMatrix contains UDI data. Check the encoded content and the issuing-agency system used; readability is also a separate verification task.
DIMDI
DIMDI was the former German Institute for Medical Documentation and Information.
The term appears in older German medical-device documents. For current activities, the responsible authorities and services of BfArM must be checked.
DI – Device Identifier
DI denotes the product-specific identification part of a UDI. UDI-DI is the corresponding more detailed designation.
Depending on the issuing agency, for example a GTIN can serve as a UDI-DI. The DI must be kept separate from the production-related PI data.
DM – Direct Marking
Direct Marking means applying the marking directly to the product.
DM is not a specific number such as a GTIN. Which identifier is used and whether direct marking is required depends on the applicable UDI context.
DPM – Direct Part Marking
Direct Part Marking means applying a marking directly to a component or product, for example by laser engraving.
An engraved serial number is not automatically a complete direct UDI marking. Data content and requirements must be checked separately.
DUNS – Data Universal Numbering System
DUNS refers to the system used by Dun & Bradstreet to identify companies and business locations.
A DUNS number is not a product identifier. Whether it is needed in a particular authority process must be checked against the current requirements.
E
EMDN – European Medical Device Nomenclature
EMDN is the European nomenclature for medical devices used for registration in EUDAMED.
The code describes a professional product category, not the individual product. EMDN is neither a UDI-DI nor automatically identical to a GMDN code.
eMDR – Electronic Medical Device Reporting
eMDR means the electronic submission of medical-device adverse-event reports to the US FDA.
Here MDR stands for Medical Device Reporting. It is not the European Medical Device Regulation and not the submission of UDI product data to GUDID.
ESG / ESG NextGen
The Electronic Submissions Gateway is an FDA-wide technical access point for electronic submissions. The current platform is called ESG NextGen.
The gateway is a transmission route to the responsible FDA processing offices, not the GUDID product database itself or a UDI data-management system.
EUDAMED – European Database on Medical Devices
EUDAMED is the European database on medical devices with several functionally different modules.
UDI/product registration, actor registration and vigilance are different tasks. The status and use of each module must be considered separately.
EURL – EU Reference Laboratory
An EU reference laboratory performs defined tasks in the IVDR context, particularly in connection with certain high-risk IVDs.
The tasks are described in Article 100 of the IVDR. A reference laboratory is not a UDI issuing agency and not a submission portal.
Evidence Pack
In this guide, a UDI Evidence Pack means the combined evidence for a submission run.
It may include the data status, approval, validation, transmission and responses. Here the term is an organisational aid, not a uniformly prescribed authority form.
F
FDA – Food and Drug Administration
The FDA is the US authority that regulates medical devices among other products.
UDI/GUDID tasks and eMDR vigilance reports use different data and processes. An authority name alone therefore does not describe a specific submission scope.
FSCA – Field Safety Corrective Action
FSCA means a field safety corrective action to reduce a product-related risk.
An FSCA, an incident report and information for users are connected but different activities. In the EU context, the MDR and IVDR are decisive.
FSN – Field Safety Notice
A Field Safety Notice is safety information for customers or users in connection with an FSCA.
It may explain the affected products, the problem and the required measures. An internal error list or an ordinary product-change notice is not automatically an FSN.
G
GDPR – General Data Protection Regulation
GDPR is the abbreviation for the EU General Data Protection Regulation.
In a data process, personal information must be distinguished from product master data. Especially in vigilance cases, technical transmission must not be planned without a privacy and authorisation concept.
GDSN – Global Data Synchronisation Network
GDSN is the network for synchronising product master data in the GS1 system.
Master-data exchange with trading partners is not automatically a regulatory UDI submission. The target system, field scope and recipient process must be clarified separately.
GHTF – Global Harmonization Task Force
GHTF was a forum of medical-device authorities and industry representatives for harmonising regulatory approaches.
Its documents remain relevant as historical foundations. The IMDRF builds on this work; GHTF and IMDRF should not be presented as the same current organisation.
GMDN – Global Medical Device Nomenclature
GMDN is an international nomenclature for describing medical-device groups.
A GMDN term is a classification or description, not an individual device identifier. It is used, among other contexts, for FDA/GUDID.
GS1
GS1 develops and maintains standards for identification, data capture and data exchange.
In the UDI process, the identifiers and carriers actually used are decisive. A GTIN record, a barcode and an authority submission remain different components.
GTIN – Global Trade Item Number
GTIN identifies a trade item in the GS1 system and can serve as a UDI-DI in the appropriate UDI context.
A GTIN is not a lot number. Different packaging levels and identification rules must be considered in data maintenance.
GUDID – Global Unique Device Identification Database
GUDID is the FDA-managed UDI product database.
The database contains DI-related product information. It is not the electronic case-reporting database for eMDR; access, data fields and submission must be handled separately.
GUDI – Global UDI SAP Add-on
GUDI is the Europe IT solution for UDI data management and authority transmission in the SAP environment.
The customer maintains the UDI data and triggers transmission from GUDI. M2M is the technical transmission route, not the entire functionality of the solution.
GSP – Global Submission Portal
GSP is the Europe IT solution for structured data imports, validation and submission processes.
The customer uploads the appropriate completed Excel template and starts transmission after successful validation. EUDAMED uses M2M / Data Exchange for this purpose.
H
Distributor under the MDR
A distributor makes a product available in the supply chain without being the manufacturer or importer.
The general distributor obligations are set out in Article 14 of the MDR. Selling a product alone does not automatically make a distributor the manufacturer or the party responsible for assigning UDI identifiers.
Primary source / further guidance ↗
WeiterlesenManufacturer under the MDR
In the MDR context, a manufacturer is the person or organisation that manufactures or has a product manufactured and markets it under its own name or trademark.
Article 10 contains central manufacturer obligations. Outsourced production or external data transmission does not automatically transfer this role to the service provider.
Primary source / further guidance ↗
WeiterlesenHIBC – Health Industry Bar Code
HIBC denotes an identification and marking system used in healthcare. HIBCC is the organisation behind it.
HIBC is not the name of one number for all products. For UDI, the applicable rules and formats of the system must be followed.
HIPAA – Health Insurance Portability and Accountability Act
HIPAA is a US law; its framework includes protection of certain health information.
Applicability depends, among other things, on the organisations and data involved. An ordinary UDI product file is not automatically a HIPAA case merely because it concerns MedTech.
HL7 – Health Level Seven
HL7 denotes an organisation and a family of standards for electronic information exchange in healthcare.
In a submission context, name the specific standard: HL7 SPL for product information and HL7 ICSR for case reports are not interchangeable.
Primary source / further guidance ↗
WeiterlesenHRI – Human Readable Interpretation
HRI is the human-readable representation of the information encoded in the data carrier.
HRI complements the machine-readable representation. It is not the same as any additional plain text on a label.
I
ICCBBA
ICCBBA manages and develops the ISBT-128 standard for identifying and marking products of human origin.
The organisation is also relevant as an issuing agency in the UDI context. Which format to use depends on the product and applicable rules.
ICSR – Individual Case Safety Report
ICSR means a structured Individual Case Safety Report.
The exact electronic standard depends on the reporting context. An FDA eMDR ICSR is not automatically a medicinal-product report under ICH E2B and is not HL7 SPL for GUDID.
IFA – Information Centre for Pharmaceutical Specialities
IFA provides, among other things, an identification system that can be used for UDI.
PZN, PPN and Basic UDI-DI are different identification concepts. The IFA system and its specific UDI application must be implemented according to the specifications.
IMDRF – International Medical Device Regulators Forum
IMDRF is an international forum of medical-device regulators promoting regulatory harmonisation and convergence.
It builds on the work of the GHTF. An IMDRF document is not automatically binding unchanged in every target market.
Importer under the MDR
In the MDR context, an importer places a product from a third country on the Union market.
Article 13 describes the general obligations. Importer, distributor and authorised representative are different roles, even if one company performs several tasks.
Primary source / further guidance ↗
WeiterlesenIssuing Agency / Issuing Entity
An issuing agency operates a system for allocating UDI identifiers.
Recognition and permitted formats must be checked for the relevant market. The issuing agency is not the product database and does not automatically submit data for the manufacturer.
IVD – In Vitro Diagnostic Medical Device
An IVD is a medical device intended for the examination outside the body of specimens derived from the human body for a medical information purpose.
In the European context, IVDs are governed by the IVDR. Risk classes A to D must not be confused with MDR classes.
IVDD – In Vitro Diagnostic Medical Devices Directive
IVDD denotes the former Directive 98/79/EC for in vitro diagnostic medical devices.
For older products, the directive context may remain relevant within applicable transitional rules. For new decisions, current IVDR requirements and transitional conditions must be checked.
IVDR – In Vitro Diagnostic Medical Devices Regulation
IVDR denotes Regulation (EU) 2017/746 on in vitro diagnostic medical devices.
Entry into force, date of application and individual transition periods are different points in time. A single year therefore does not explain a product’s complete regulatory status.
IOL – Intraocular Lens
An intraocular lens is a lens implanted in the eye.
An IOL and a contact lens are not the same. Rules for one optical product group must not be transferred to another solely because similar terms are used.
J
JSON – JavaScript Object Notation
JSON is a text-based format for the structured representation and exchange of data.
A JSON file says nothing by itself about substantive correctness or possible authority acceptance. The data model, interface contract and validation remain necessary.
K
Kit under the IVDR
In the IVDR context, a kit is a group of components packaged together for a specific in vitro examination or part of one.
The term from Article 2 of the IVDR must not be equated without checking it with an MDR treatment pack or an FDA convenience kit.
Kit / Convenience Kit in the FDA context
The FDA uses Convenience Kit for a specific combination of different medical devices supplied together for the user.
Not every product combination is automatically a Convenience Kit in the UDI context. The FDA definition and conditions of use must be checked separately.
Clinical Investigation
A clinical investigation systematically examines the safety or performance of a medical device with human participants.
It is a different task from UDI data validation. In the European legal framework, the provisions in Chapter VI of the MDR are relevant, among others.
L
Storage and handling conditions
These details describe the conditions under which a product should be stored, transported or handled.
In a UDI data model, type, unit, limits and free text can be separate fields. The data must match the product documentation.
Latex
Latex or natural-rubber latex may be relevant as a material declaration and in connection with allergy risks.
Whether and how a declaration is required on a label or in a database depends on the material, product and target market. Do not treat every medical device as latex-containing or subject to identical declaration duties.
Legacy Device
In the EU context, a Legacy Device is a product still placed on the market under the former directive-based law in accordance with the applicable transitional provisions.
It does not simply mean every old product. In EUDAMED, special identification concepts such as EUDAMED DI and EUDAMED ID may apply to Legacy Devices.
Legislation
Legislation denotes the legal framework of a subject area.
In the European medical-device environment, the MDR and IVDR and, where applicable, former directives and transitional rules are particularly relevant. An authority guide, a law and a technical specification are different document types.
M
Master UDI-DI
Master UDI-DI is a particular UDI allocation concept for certain highly individualised products, including contact lenses and certain spectacle products.
It is not a general superordinate identifier for every product portfolio and is not equivalent to Basic UDI-DI. The product scope and introduction follow the applicable EU rules.
MDCG – Medical Device Coordination Group
MDCG is the Medical Device Coordination Group in the European regulatory framework.
Its guidance supports consistent application. An MDCG guidance document must be distinguished from the underlying regulation; the document version and scope are relevant.
MDD – Medical Devices Directive
MDD denotes the former European Medical Devices Directive 93/42/EEC.
The term remains relevant in Legacy and transitional contexts. An old MDD reference must not be used as blanket evidence of current MDR conformity.
MDR – Medical Device Regulation
In the European context, MDR denotes Regulation (EU) 2017/745 on medical devices.
In US documents, MDR can instead mean Medical Device Reporting. The same abbreviation therefore refers to different regulatory tasks.
MDSAP – Medical Device Single Audit Program
MDSAP enables one audit of a medical-device manufacturer against the relevant requirements of several participating regulators.
The programme does not automatically replace every product registration or regulatory obligation. Its scope and the roles of participating authorities must be checked against the current programme documents.
MIR – Manufacturer Incident Report
An MIR is used in the European vigilance context for a structured report of a serious incident by the manufacturer.
An MIR report is not a UDI product registration. Case assessment, the responsible reporting route and data transmission must be distinguished.
M2M – Machine-to-Machine
M2M means technical data exchange between systems.
At Europe IT, GSP and GUDI can use the M2M route in the respective authority process. The solutions, data source and operating model remain distinct; M2M is not automatically an individual interface-building project at the customer.
MTR – Manufacturer’s Trend Report
A trend report concerns a relevant change in the frequency or severity of certain events under the applicable vigilance framework.
It must be distinguished from an individual incident report. The reporting criteria arise from the MDR or IVDR and must not be reduced to any arbitrary change in numbers.
N
NANDO
NANDO denotes the European information system for Notified Bodies and their respective scope of designation.
NANDO is not a Notified Body such as a testing organisation. When selecting a body, check its specific designation and the product areas it covers.
NDC – National Drug Code
NDC is a US identifier used in the pharmaceutical sector.
It is not a UDI-DI for medical devices. Drug and medical-device identifiers must not be equated simply because both occur in FDA processes.
Notified Body
A Notified Body is an organisation designated for specific conformity-assessment tasks.
Its activities are limited to the designation concerned. It is neither the responsible supervisory authority nor a UDI issuing agency or submission platform.
O
OCR – Optical Character Recognition
OCR means the machine recognition of text in images or scanned documents.
For labels, OCR can support the capture of plain text. It does not replace decoding a barcode or the substantive review of the captured information.
Owner / Operator
Owner/Operator denotes the owner or operator of an establishment in the FDA registration context.
This organisational role must not be equated without checking it with the UDI labeler. The party responsible for UDI labelling is determined under the applicable FDA rules.
Off-label use
Off-label use means use outside the application intended by the manufacturer or specified in the applicable regulatory framework.
The term should not be confused with changing a published UDI record. Questions about a specific use require their own professional and regulatory assessment.
OEM – Original Equipment Manufacturer
OEM means an Original Equipment Manufacturer, for example a producer of components or products offered under another brand.
The commercial designation alone does not determine who is the regulatory manufacturer. This role follows from the specific product and marketing context.
Obsolescence Management
Obsolescence Management addresses risks arising from components, materials and technologies that are being phased out or are no longer available.
It is not a blanket synonym for Regulatory Compliance. Changes may require an assessment of their impact on product data, labelling and identifiers.
P
PPN – Pharmacy Product Number
Pharmacy Product Number is an identifier in the IFA system that can, among other things, incorporate national article-number systems.
PPN and a national PZN are not identical. For UDI use, structure, allocation and coding must follow the applicable IFA specification.
PI – Production Identifier
PI denotes production-related components of a UDI, such as lot, serial number, manufacturing date or expiry date, where applicable.
PI types in the master record and concrete PI values on the product must be distinguished. Not every UDI database stores the value of every individual unit produced.
PRA Code – Product Registration Agency Code
The PRA code is part of the IFA identification concept for distinguishing numbering systems.
It is not, by itself, the complete Pharmacy Product Number. Use the IFA specification for the concrete construction and check-digit logic.
Primary source / further guidance ↗
Procedure Pack
In the MDR context, a procedure pack is a group of products packaged and placed on the market together for a specific medical purpose.
The term must be distinguished from an IVDR kit and from any packaging level. The data model and responsibilities must fit the type of combination.
PSR – Periodic Summary Report
A PSR is a periodic summary report on certain similar incidents.
It does not replace individual reports at will. In the European vigilance context, the procedure requires corresponding coordination with the competent authority.
PSRP – PSR Periodic Analysis Update
In the EUDAMED vigilance context, PSRP denotes a periodic analysis update to an existing PSR.
The authority help area describes the link to an active PSR. The availability of a technical instruction alone does not prove that a module can be used productively.
PSUR – Periodic Safety Update Report
A PSUR summarises safety information from post-market surveillance.
Do not mix the classes: under the MDR, the PSUR context particularly concerns IIa, IIb and III; under the IVDR, classes C and D. Scope and update details must be checked separately.
PZN – German Pharmaceutical Product Number
PZN is a German article identifier used in pharmaceutical and pharmacy-related commerce.
A PZN is not automatically the same identifier as a PPN or a UDI-DI. The relationships must be represented clearly in the coding system used.
Q
QMS – Quality Management System
A quality management system organises processes, responsibilities and evidence for meeting quality requirements.
UDI data maintenance can be integrated into the QMS. A software product or completed template alone is not a complete quality management system.
QM – Quality Management
Quality management includes the quality-oriented direction, planning, control and improvement of activities.
In a UDI process, this may include responsibilities, data quality and correction workflows. QM is broader than a single technical validation check.
QR Code
A QR code is a two-dimensional data carrier that can encode, for example, text or a URL.
A QR code is not automatically a UDI carrier and is not the same as a DataMatrix. The permitted use and syntax must fit the purpose.
QS / QSR / QMSR
QS generally means Quality System. In the FDA medical-device context, distinguish the former Quality System Regulation from the current QMSR.
The FDA QMSR has applied since 2 February 2026. Do not carry older references to the former QS framework over as unchanged current requirements.
R
REST – Representational State Transfer
REST is an architectural style for distributed systems and web interfaces.
REST is not an authority format such as HL7 SPL and is not evidence that GUDID accepts arbitrary REST calls. An internal API and the subsequent authority channel can have different technical roles.
RFID – Radio-Frequency Identification
RFID enables identification using radio communication between a transponder and a reader.
RFID and optical barcodes are different capture technologies. Whether a particular carrier is suitable for the intended UDI use must be assessed independently of the radio method itself.
Resubmission
Resubmission means submitting again, for example after correcting a response.
The new run should be documented with the changed data status and a reference to the original result. For GSP/GUDI data errors, substantive correction remains the customer’s responsibility.
S
SOX – Sarbanes-Oxley Act
SOX denotes a US legal framework for corporate and financial reporting.
SOX is not a UDI regulation. Financial-control requirements must not be described broadly as regulatory obligations for every UDI software product.
Software Version
The software version identifies a development or release state of software.
For software as a medical device, changes can affect the UDI-DI or UDI-PI. Not every commercial software product is automatically a medical device; the applicable UDI rules must be checked.
Primary source / further guidance ↗
WeiterlesenCustom-Made Device
A custom-made device is made for the individual conditions and needs of a particular patient following an appropriate written prescription.
Not every adapted off-the-shelf product thereby becomes a custom-made device. Product type and regulatory definition are decisive, not merely the manufacturing technology.
Special Device Type
Special Device Type is a data-model-specific classification for special product types.
The available values and their meaning depend on the regulatory context and target system. A drop-down list does not replace a professional assessment of the product.
WeiterlesenSRN – Single Registration Number
SRN is a registration identifier for economic operators in the European EUDAMED context.
It is not a UDI-DI or a Basic UDI-DI. In a submission, the actor identifier and product identifiers must be assigned to the correct fields.
Sterile
Sterile describes the state of freedom from viable microorganisms within the relevant sterility concept.
In the product record, for example, sterile supply and sterilisation before use must be distinguished. A simple data entry is not evidence that the sterilisation process has been validated.
swissdamed
swissdamed is the Swiss database for economic operators and medical devices operated by Swissmedic.
It is modelled on EUDAMED but is a separate system. An existing EUDAMED registration does not automatically mean that a Swiss product registration has been completed.
Submission
Submission means submitting information to an authority or another target system.
Data preparation, validation, transmission and the processing result are separate steps. A successful pre-check alone is not a confirmed authority acceptance.
T
TGA – Therapeutic Goods Administration
TGA is the Australian regulatory authority for therapeutic goods, including medical devices.
For UDI product data, it operates AusUDID. The authority name, UDI database and specific regulatory task should be named separately in project descriptions.
Third Party
A third party is an external participant that performs defined tasks in the agreed process.
Europe IT can support the technical UDI transmission process as a provider. Commissioning a third party does not automatically make it the legal manufacturer, sponsor or holder of all professional responsibility.
Type of Multi-Component Device
This term denotes a data-model-specific classification of multi-component products.
The selection depends on the product composition, MDR/IVDR context and target system. A multi-component product is not simply any package containing several articles.
WeiterlesenU
UDI – Unique Device Identification
UDI denotes a system for uniquely identifying medical devices.
Where applicable, a UDI consists of a product-specific and a production-specific part. Labelling, internal master data and authority information must fit together, but remain different tasks.
UDI-DI and Basic UDI-DI
UDI-DI and Basic UDI-DI have different roles in the European data model: product identifier and overarching regulatory grouping.
Several UDI-DIs can be assigned to a suitable Basic UDI-DI. The inventory illustration below shows the principle, but is not a complete specification or a valid number template.
Primary source / further guidance ↗
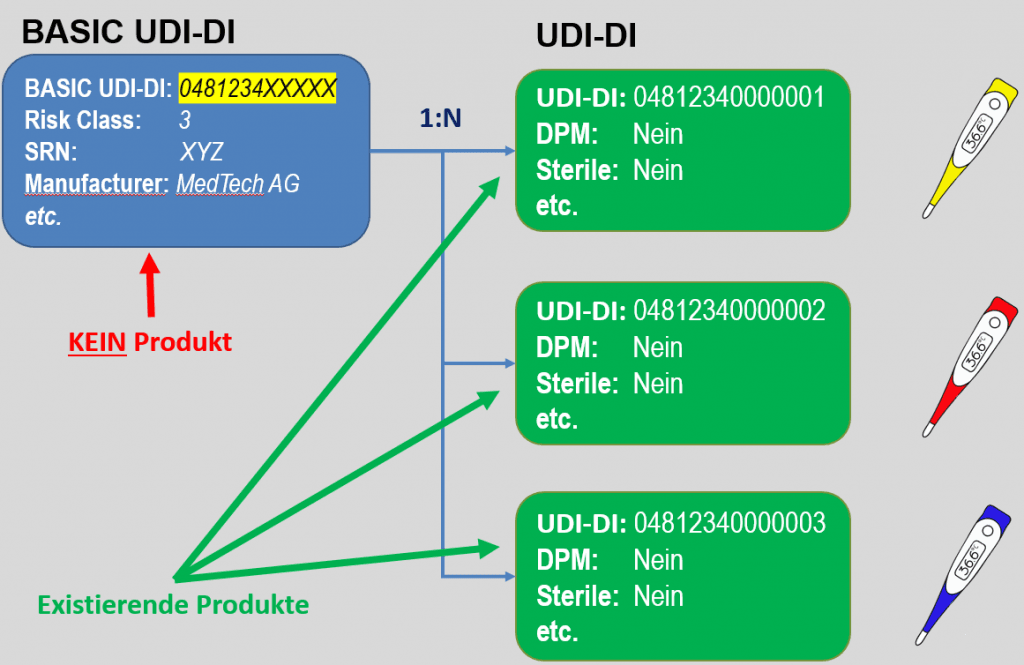
EUDAMED Basic UDI-DI and UDI-DI
Existing illustration by Europe IT: schematic assignment only; the example identifiers have not been checked. The Basic UDI-DI and product record remain separate.
Unit of Use DI
A Unit of Use DI identifies the unit of use in the relevant UDI context, especially where not every individual unit of use carries its own UDI carrier.
It may become relevant, for example, for a pack containing multiple units. Its role must be distinguished from the packaging identifier and the production-related serial number.
V
Validation
Validation must be explained in its specific context. UDI data validation checks data against the rules and structures represented in the process.
This is not automatically the same as validating software for its intended use. A data check without errors is also not a comprehensive regulatory product release.
Vigilance
Vigilance is the system for handling and reporting safety-relevant incidents and measures in the medical-device context.
It is part of post-market product-safety activities, but must not be equated with all market-surveillance activities or with UDI registration.
Verification
Verification checks whether specified requirements have been met.
In a data or labelling process, this may include field rules, barcode content or readability. The subject and criteria of the check must be named; the word alone does not describe a complete compliance assessment.
Volume Production
Volume production means manufacturing larger quantities.
A high unit count is not automatically a high number of different UDI-DIs. Unit quantities, product variants and authority records must not be confused when planning the process.
Seller
Seller is a general commercial designation.
The applicable regulatory obligations depend on the actual role, such as manufacturer, importer or distributor. A seller therefore does not always have to assign new UDI identifiers itself.
VDP – Variable Data Printing
Variable Data Printing means printing changing content within a print process, such as a lot or serial number on labels.
Static product data and variable production data must be combined correctly. Printing, data content and checking the marking are separate tasks.
W
Warnings and Precautions
Warnings and precautions provide information about relevant risks and the safe handling of a product.
In labelling, instructions for use and product data, the information must fit the intended product. Generic warning text is not a substitute for the actual product documentation.
WebTrader
WebTrader was the former FDA web interface for electronic submissions.
According to the FDA, it was replaced by the Unified Submission Portal of ESG NextGen on 14 April 2025. Old WebTrader instructions are therefore not unchanged current operating instructions.
WHO – World Health Organization
The World Health Organization is the health organisation of the United Nations.
Among other things, it publishes international health information and guidance. A WHO publication is not automatically a national UDI submission requirement.
X
XML – Extensible Markup Language
XML is a markup language for the structured representation of data.
Generating XML, checking a schema and transmitting the file are different tasks. A file can be valid XML and still not match the required authority format.
XSD – XML Schema Definition
An XSD describes permitted structures and data types of an XML document.
A successful XSD check does not prove that all substantive dependencies, permissions or authority rules are met. Schema validation and business validation must be considered separately.
Y
Yes / No – Controlled Data Values
Yes and No are typical selection values in structured records.
If a template or interface requires specific codes, they must not be replaced arbitrarily by “Yes”, “No”, “1” or free text. The concrete code list, not everyday language, is decisive.
Z
ZIP
ZIP is a file format for compressing and bundling files.
A ZIP archive is not automatically a complete Evidence Pack. Content, linkage to the data status and submission, and controlled storage remain separate tasks.
Approval / Registration Number
An approval or registration number refers to a procedure or record in the relevant regulatory system.
It is not automatically part of a UDI and is not equivalent to a UDI-DI. The data model must represent the product identifier and regulatory reference separately.
Certification
Certification means confirmation of specified requirements by an authorised body within a defined scope.
A certificate applies to its specific subject and scope. UDI transmission or a technical acceptance status is not a general product or company certification.
SSCP – Summary of Safety and Clinical Performance
SSCP is a Summary of Safety and Clinical Performance in the MDR context.
Article 32 defines its scope and content. It is not required in the same way for every medical device and is not the same as a PSUR.
Competent Authorities
Competent authorities perform the tasks for medical devices assigned to them by the applicable legal framework.
Authority, Notified Body, UDI issuing agency and technical service provider must not be presented as interchangeable roles. See also CA and Notified Body.
Intended Purpose
The intended purpose describes the use of a product intended by the manufacturer.
It is essential for regulatory classification and many data decisions. Changes to advertising, labelling or instructions for use must therefore also be assessed in their professional context.
From term to next work step
UDI guide · UDI data transmission · Submission evidence · Ask a UDI question
Terms explain the context. Whether a particular requirement applies to your product or organisation must be clarified based on the target market, your role and the current documentation.
Editorial source check: 4 September 2026. Short definitions and explanations are based on the existing Europe IT glossary and were supplemented and corrected using primary sources. Linked complete regulations and download packages have not been fully implemented or functionally tested.
